Recombinant Human Myosin-7 (MYH7), partial

Recombinant Human Myosin-7 (MYH7), partial — Myosins are actin-based motor molecules with ATPase activity essential for muscle contraction. Purity >90%.

SKU: BCRECP-001207 Category:

Product Specifications

Product SkuBCRECP-001207
Product DescriptionRecombinant Human Myosin-7 (MYH7) Protein is expressed from E.coli with N-terminal 10xHis-tagged. It contains 1-109aa. [Accession | P12883].
Uniprot No.P12883
Gene NamesMYH7
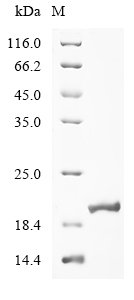
PurityGreater than 90% as determined by SDS-PAGE.
Expression SystemE.coli
Expression Region1-109aa
SpeciesHomo sapiens (Human)
Tag InfoN-terminal 10xHis-tagged
Molecular weight18.5kDa
ActivityPlease contact us to obtain bioactivity data.
BufferIf the delivery form is liquid, the default storage buffer is Tris/PBS-based buffer, 5%-50% glycerol. If the delivery form is lyophilized powder, the buffer before lyophilization is Tris/PBS-based buffer, 6% Trehalose.
StorageStore at -20°C/-80°C upon receipt, aliquoting is necessary for mutiple use. Avoid repeated freeze-thaw cycles.
Research AreasSignal Transduction

Function

Myosins are actin-based motor molecules with ATPase activity essential for muscle contraction. Forms regular bipolar thick filaments that, together with actin thin filaments, constitute the fundamental contractile unit of skeletal and cardiac muscle.

Biological Context

Subcellular Location: Cytoplasm, myofibril; Cytoplasm, myofibril, sarcomere
Tissue Specificity: Both wild type and variant Gln-403 are detected in skeletal muscle (at protein level)
Disease Association: Cardiomyopathy, familial hypertrophic, 1 (CMH1) : A hereditary heart disorder characterized by ventricular hypertrophy, which is usually asymmetric and often involves the interventricular septum. The symptoms include dyspnea, syncope, collapse, palpitations, and chest pain. They can be readily provoked by exercise. The disorder has inter- and intrafamilial variability ranging from benign to malignant forms with high risk of cardiac failure and sudden cardiac death. [The disease is caused by variants affecting the gene represented in this entry] | Congenital myopathy 7A, myosin storage, autosomal dominant (CMYO7A) : A skeletal muscle disorder characterized by prominent axial and proximal weakening, spinal stiffness, severe scoliosis, with or without respiratory and cardiac involvement. The age at symptom onset can range from early childhood to late adulthood, and disease severity ranges from asymptomatic to severe muscular weakness and respiratory insufficiency. Histopathological examination shows variable findings including subsarcolemmal hyaline bodies in type 1 fibers. [The disease is caused by variants affecting the gene represented in this entry] | Cardiomyopathy, dilated, 1S (CMD1S) : A disorder characterized by ventricular dilation and impaired systolic function, resulting in congestive heart failure and arrhythmia. Patients are at risk of premature death. [The disease is caused by variants affecting the gene represented in this entry] | Myopathy, distal, 1 (MPD1) : A muscular disorder characterized by early-onset selective weakness of the great toe and ankle dorsiflexors, followed by weakness of the finger extensors. Mild proximal weakness occasionally develops years later after the onset of the disease. [The disease is caused by variants affecting the gene represented in this entry] | Congenital myopathy 7B, myosin storage, autosomal recessive (CMYO7B) : A skeletal muscle disorder characterized by the onset of scapuloperoneal muscle weakness in early childhood or young adulthood. Affected individuals have difficulty walking, steppage gait, and scapular winging due to shoulder girdle involvement. The severity and progression of the disorder is highly variable. Most patients develop respiratory insufficiency and restrictive lung disease. Some develop hypertrophic cardiomyopathy. Histopathological examination shows variable findings including subsarcolemmal hyaline bodies in type 1 fibers. [The disease is caused by variants affecting the gene represented in this entry] | Left ventricular non-compaction 5 (LVNC5) : A form of left ventricular non-compaction, a cardiomyopathy due to myocardial morphogenesis arrest and characterized by a hypertrophic left ventricle, a severely thickened 2-layered myocardium, numerous prominent trabeculations, deep intertrabecular recesses, and poor systolic function. Clinical manifestations are variable. Some affected individuals experience no symptoms at all, others develop heart failure. In some cases, left ventricular non-compaction is associated with other congenital heart anomalies. LVNC5 is an autosomal dominant condition. [The disease is caused by variants affecting distinct genetic loci, including the gene represented in this entry]

Product Specifications

Recombinant Human Myosin-7 (MYH7), partial is a recombinant protein from Homo sapiens (Human), expressed in E.coli, covering amino acids 1-109aa, with N-terminal 10xHis-tagged tag, molecular weight 18.5kDa, purity Greater than 90% as determined by SDS-PAGE.. Suitable for ELISA and Western Blot applications.
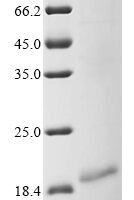
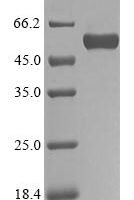
Recommended applications depend on the target, product format, and product-specific validation data. Refer to the product datasheet or Certificate of Analysis (CoA), or contact BioCrest Sci for application guidance.

Biological Activity: Please contact us to obtain bioactivity data.

Storage: Store at -20°C/-80°C upon receipt, aliquoting is necessary for mutiple use. Avoid repeated freeze-thaw cycles.

Shipping & Lead Time: Availability, shipping conditions, and estimated lead time are confirmed in the formal quotation or order confirmation based on the product format, destination, and order requirements.

Protein Biology

Function

Myosins are actin-based motor molecules with ATPase activity essential for muscle contraction. Forms regular bipolar thick filaments that, together with actin thin filaments, constitute the fundamental contractile unit of skeletal and cardiac muscle

Subcellular Location

Cytoplasm, myofibril; Cytoplasm, myofibril, sarcomere

Disease Association

Cardiomyopathy, familial hypertrophic, 1 (CMH1) : A hereditary heart disorder characterized by ventricular hypertrophy, which is usually asymmetric and often involves the interventricular septum. The symptoms include dyspnea, syncope, collapse, palpitations, and chest pain. They can be readily provoked by exercise. The disorder has inter- and intrafamilial variability ranging from benign to malignant forms with high risk of cardiac failure and sudden cardiac death. [The disease is caused by variants affecting the gene represented in this entry] | Congenital myopathy 7A, myosin storage, autosomal dominant (CMYO7A) : A skeletal muscle disorder characterized by prominent axial and proximal weakening, spinal stiffness, severe scoliosis, with or without respiratory and cardiac involvement. The age at symptom onset can range from early childhood to late adulthood, and disease severity ranges from asymptomatic to severe muscular weakness and respiratory insufficiency. Histopathological examination shows variable findings including subsarcolemmal hyaline bodies in type 1 fibers. [The disease is caused by variants affecting the gene represented in this entry] | Cardiomyopathy, dilated, 1S (CMD1S) : A disorder characterized by ventricular dilation and impaired systolic function, resulting in congestive heart failure and arrhythmia. Patients are at risk of premature death. [The disease is caused by variants affecting the gene represented in this entry] | Myopathy, distal, 1 (MPD1) : A muscular disorder characterized by early-onset selective weakness of the great toe and ankle dorsiflexors, followed by weakness of the finger extensors. Mild proximal weakness occasionally develops years later after the onset of the disease. [The disease is caused by variants affecting the gene represented in this entry] | Congenital myopathy 7B, myosin storage, autosomal recessive (CMYO7B) : A skeletal muscle disorder characterized by the onset of scapuloperoneal muscle weakness in early childhood or young adulthood. Affected individuals have difficulty walking, steppage gait, and scapular winging due to shoulder girdle involvement. The severity and progression of the disorder is highly variable. Most patients develop respiratory insufficiency and restrictive lung disease. Some develop hypertrophic cardiomyopathy. Histopathological examination shows variable findings including subsarcolemmal hyaline bodies in type 1 fibers. [The disease is caused by variants affecting the gene represented in this entry] | Left ventricular non-compaction 5 (LVNC5) : A form of left ventricular non-compaction, a cardiomyopathy due to myocardial morphogenesis arrest and characterized by a hypertrophic left ventricle, a severely thickened 2-layered myocardium, numerous prominent trabeculations, deep intertrabecular recesses, and poor systolic function. Clinical manifestations are variable. Some affected individuals experience no symptoms at all, others develop heart failure. In some cases, left ventricular non-compaction is associated with other congenital heart anomalies. LVNC5 is an autosomal dominant condition. [The disease is caused by variants affecting distinct genetic loci, including the gene represented in this entry]

Tissue Specificity

Both wild type and variant Gln-403 are detected in skeletal muscle (at protein level)

Subunit

Muscle myosin is a hexameric protein that consists of 2 heavy chain subunits (MHC), 2 alkali light chain subunits (MLC) and 2 regulatory light chain subunits (MLC-2). Interacts with ECPAS (PubMed:20682791). Interacts (via C-terminus) with LRRC39 (PubMed:20847312)

Gene: MYH7  |  Organism: Homo sapiens  |  Synonyms: Myosin heavy chain 7; Myosin heavy chain slow isoform; Myosin heavy chain, cardiac muscle beta isoform
Key Publications

Frequently Asked Questions

How do I order or inquire about this product?

Submit the Online Inquiry form with your required quantity and specifications, or email sales@biocrestsci.com. Available sizes, pricing, stock status, documentation, and estimated lead time are confirmed in the formal quotation.

What is the shipping and delivery time?

Availability, dispatch timing, shipping conditions, and delivery estimates depend on the product format, destination, and order requirements. These details are confirmed in the formal quotation or order confirmation.

How should this product be stored?

Follow the storage and handling instructions in the product-specific datasheet and Certificate of Analysis (CoA). Requirements may vary by product form and formulation; contact BioCrest Sci if product-specific guidance is needed.

What quality controls are performed on your products?

Quality-control data such as identity, purity, endotoxin level, or biological activity are provided only when specified on this product page, the datasheet, or Certificate of Analysis (CoA). Biological activity is not implied unless explicitly stated.

Shopping Cart
Scroll to Top