Recombinant Human Alpha-L-iduronidase (IDUA)

Recombinant Human Alpha-L-iduronidase (IDUA) — recombinant protein from Homo sapiens (Human), expressed in E.coli. Purity >90%.

SKU: BCRECP-000884 Category:

Product Specifications

Product SkuBCRECP-000884
Product DescriptionRecombinant Human Alpha-L-iduronidase (IDUA) Protein is expressed from E.coli with N-terminal 6xHis-tagged. It contains 28-653aa. [Accession | P35475].
Uniprot No.P35475
Gene NamesIDUA
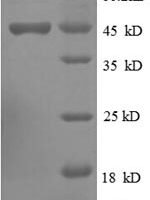
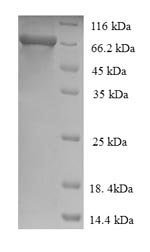
PurityGreater than 90% as determined by SDS-PAGE.
Expression SystemE.coli
Expression Region28-653aa
SpeciesHomo sapiens (Human)
Tag InfoN-terminal 6xHis-tagged
Molecular weight73.9kDa
ActivityPlease contact us to obtain bioactivity data.
BufferIf the delivery form is liquid, the default storage buffer is Tris/PBS-based buffer, 5%-50% glycerol. If the delivery form is lyophilized powder, the buffer before lyophilization is Tris/PBS-based buffer, 6% Trehalose.
StorageStore at -20°C/-80°C upon receipt, aliquoting is necessary for mutiple use. Avoid repeated freeze-thaw cycles.
Research AreasMetabolism

Function

Lysosomal hydrolase responsible for the degradation of the glycosaminoglycans heparan sulfate and dermatan sulfate.

Biological Context

Subcellular Location: Lysosome
Tissue Specificity: Ubiquitous
Disease Association: Mucopolysaccharidosis 1H (MPS1H) : A severe form of mucopolysaccharidosis type 1, a rare lysosomal storage disease characterized by progressive physical deterioration with urinary excretion of dermatan sulfate and heparan sulfate. Patients with MPS1H usually present, within the first year of life, a combination of hepatosplenomegaly, skeletal deformities, corneal clouding and severe intellectual disability. Obstructive airways disease, respiratory infection and cardiac complications usually result in death before 10 years of age. [The disease is caused by variants affecting the gene represented in this entry] | Mucopolysaccharidosis 1H/S (MPS1H/S) : A form of mucopolysaccharidosis type 1, a rare lysosomal storage disease characterized by progressive physical deterioration with urinary excretion of dermatan sulfate and heparan sulfate. MPS1H/S represents an intermediate phenotype of the MPS1 clinical spectrum. It is characterized by relatively little neurological involvement, but most of the somatic symptoms described for severe MPS1 develop in the early to mid-teens, causing considerable loss of mobility. [The disease is caused by variants affecting the gene represented in this entry] | Mucopolysaccharidosis 1S (MPS1S) : A mild form of mucopolysaccharidosis type 1, a rare lysosomal storage disease characterized by progressive physical deterioration with urinary excretion of dermatan sulfate and heparan sulfate. Patients with MPS1S may have little or no neurological involvement, normal stature and life span, but present development of joints stiffness, mild hepatosplenomegaly, aortic valve disease and corneal clouding. [The disease is caused by variants affecting the gene represented in this entry]

Product Specifications

Recombinant Human Alpha-L-iduronidase (IDUA) is a recombinant protein from Homo sapiens (Human), expressed in E.coli, covering amino acids 28-653aa, with N-terminal 6xHis-tagged tag, molecular weight 73.9kDa, purity Greater than 90% as determined by SDS-PAGE.. Suitable for ELISA and Western Blot applications.
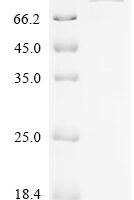
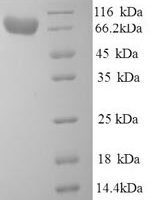
Recommended applications depend on the target, product format, and product-specific validation data. Refer to the product datasheet or Certificate of Analysis (CoA), or contact BioCrest Sci for application guidance.

Biological Activity: Please contact us to obtain bioactivity data.

Storage: Store at -20°C/-80°C upon receipt, aliquoting is necessary for mutiple use. Avoid repeated freeze-thaw cycles.

Shipping & Lead Time: Availability, shipping conditions, and estimated lead time are confirmed in the formal quotation or order confirmation based on the product format, destination, and order requirements.

Protein Biology

Function

Lysosomal hydrolase responsible for the degradation of the glycosaminoglycans heparan sulfate and dermatan sulfate

Subcellular Location

Lysosome

Disease Association

Mucopolysaccharidosis 1H (MPS1H) : A severe form of mucopolysaccharidosis type 1, a rare lysosomal storage disease characterized by progressive physical deterioration with urinary excretion of dermatan sulfate and heparan sulfate. Patients with MPS1H usually present, within the first year of life, a combination of hepatosplenomegaly, skeletal deformities, corneal clouding and severe intellectual disability. Obstructive airways disease, respiratory infection and cardiac complications usually result in death before 10 years of age. [The disease is caused by variants affecting the gene represented in this entry] | Mucopolysaccharidosis 1H/S (MPS1H/S) : A form of mucopolysaccharidosis type 1, a rare lysosomal storage disease characterized by progressive physical deterioration with urinary excretion of dermatan sulfate and heparan sulfate. MPS1H/S represents an intermediate phenotype of the MPS1 clinical spectrum. It is characterized by relatively little neurological involvement, but most of the somatic symptoms described for severe MPS1 develop in the early to mid-teens, causing considerable loss of mobility. [The disease is caused by variants affecting the gene represented in this entry] | Mucopolysaccharidosis 1S (MPS1S) : A mild form of mucopolysaccharidosis type 1, a rare lysosomal storage disease characterized by progressive physical deterioration with urinary excretion of dermatan sulfate and heparan sulfate. Patients with MPS1S may have little or no neurological involvement, normal stature and life span, but present development of joints stiffness, mild hepatosplenomegaly, aortic valve disease and corneal clouding. [The disease is caused by variants affecting the gene represented in this entry]

Tissue Specificity

Ubiquitous

Subunit

Monomer

Gene: IDUA  |  Organism: Homo sapiens
Key Publications

Frequently Asked Questions

How do I order or inquire about this product?

Submit the Online Inquiry form with your required quantity and specifications, or email sales@biocrestsci.com. Available sizes, pricing, stock status, documentation, and estimated lead time are confirmed in the formal quotation.

What is the shipping and delivery time?

Availability, dispatch timing, shipping conditions, and delivery estimates depend on the product format, destination, and order requirements. These details are confirmed in the formal quotation or order confirmation.

How should this product be stored?

Follow the storage and handling instructions in the product-specific datasheet and Certificate of Analysis (CoA). Requirements may vary by product form and formulation; contact BioCrest Sci if product-specific guidance is needed.

What quality controls are performed on your products?

Quality-control data such as identity, purity, endotoxin level, or biological activity are provided only when specified on this product page, the datasheet, or Certificate of Analysis (CoA). Biological activity is not implied unless explicitly stated.

Shopping Cart
Scroll to Top