Recombinant Human Collagen alpha-1 (I) chain (COL1A1), partial

Recombinant Human Collagen alpha-1 (I) chain (COL1A1), partial — recombinant protein from Homo sapiens (Human), expressed in Yeast. Purity >90%.

SKU: BCRECP-000403 Category:

Product Specifications

Product SkuBCRECP-000403
Product DescriptionRecombinant Human Collagen alpha-1 (I) chain (COL1A1) Protein is expressed from Yeast with Confidential. It contains Confidential. [Accession | P02452].
Uniprot No.P02452
Gene NamesCOL1A1
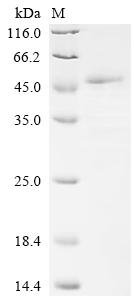
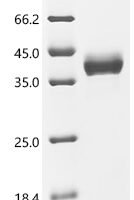
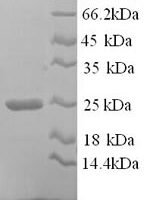
PurityGreater than 90% as determined by SDS-PAGE.
Expression SystemYeast
Expression RegionConfidential
SpeciesHomo sapiens (Human)
Tag InfoConfidential
Molecular weight47.0kDa
ActivityPlease contact us to obtain bioactivity data.
BufferIf the delivery form is liquid, the default storage buffer is Tris/PBS-based buffer, 5%-50% glycerol. If the delivery form is lyophilized powder, the buffer before lyophilization is Tris/PBS-based buffer, 6% Trehalose.
StorageStore at -20°C/-80°C upon receipt, aliquoting is necessary for mutiple use. Avoid repeated freeze-thaw cycles.
Research AreasOthers

Function

Type I collagen is a member of group I collagen (fibrillar forming collagen).

Biological Context

Subcellular Location: Secreted, extracellular space, extracellular matrix
Tissue Specificity: Forms the fibrils of tendon, ligaments and bones. In bones the fibrils are mineralized with calcium hydroxyapatite
Disease Association: Caffey disease (CAFYD) : An autosomal dominant disorder characterized by an infantile episode of massive subperiosteal new bone formation that typically involves the diaphyses of the long bones, mandible, and clavicles. The involved bones may also appear inflamed, with painful swelling and systemic fever often accompanying the illness. The bone changes usually begin before 5 months of age and resolve before 2 years of age. [The disease is caused by variants affecting the gene represented in this entry] | Ehlers-Danlos syndrome, classic type, 1 (EDSCL1) : A form of Ehlers-Danlos syndrome, a group of connective tissue disorders characterized by skin hyperextensibility, articular hypermobility, and tissue fragility. The main features of classic Ehlers-Danlos syndrome are joint hypermobility and dislocation, and fragile, bruisable skin. EDSCL1 inheritance is autosomal dominant. [The disease may be caused by variants affecting the gene represented in this entry] | Ehlers-Danlos syndrome, arthrochalasia type, 1 (EDSARTH1) : A form of Ehlers-Danlos syndrome, a connective tissue disorder characterized by hyperextensible skin, atrophic cutaneous scars due to tissue fragility and joint hyperlaxity. EDSARTH1 is an autosomal dominant form characterized by frequent congenital hip dislocation and extreme joint laxity with recurrent joint subluxations and minimal skin involvement. [The disease is caused by variants affecting the gene represented in this entry] | Osteogenesis imperfecta 1 (OI1) : An autosomal dominant form of osteogenesis imperfecta, a disorder of bone formation characterized by bone low bone mass, bone fragility and susceptibility to fractures after minimal trauma. Disease severity ranges from very mild forms without fractures to intrauterine fractures and perinatal lethality. Extraskeletal manifestations, which affect a variable number of patients, are dentinogenesis imperfecta, hearing loss, and blue sclerae. OI1 is a non-deforming form with normal height or mild short stature, and no dentinogenesis imperfecta. [The disease is caused by variants affecting the gene represented in this entry] | Osteogenesis imperfecta 2 (OI2) : An autosomal dominant form of osteogenesis imperfecta, a disorder of bone formation characterized by low bone mass, bone fragility and susceptibility to fractures after minimal trauma. Disease severity ranges from very mild forms without fractures to intrauterine fractures and perinatal lethality. Extraskeletal manifestations, which affect a variable number of patients, are dentinogenesis imperfecta, hearing loss, and blue sclerae. OI2 is characterized by bone fragility, with many perinatal fractures, severe bowing of long bones, undermineralization, and death in the perinatal period due to respiratory insufficiency. [The disease is caused by variants affecting the gene represented in this entry] | Osteogenesis imperfecta 3 (OI3) : An autosomal dominant form of osteogenesis imperfecta, a disorder of bone formation characterized by low bone mass, bone fragility and susceptibility to fractures after minimal trauma. Disease severity ranges from very mild forms without fractures to intrauterine fractures and perinatal lethality. Extraskeletal manifestations, which affect a variable number of patients, are dentinogenesis imperfecta, hearing loss, and blue sclerae. OI3 is characterized by progressively deforming bones, very short stature, a triangular face, severe scoliosis, grayish sclera and dentinogenesis imperfecta. [The disease is caused by variants affecting the gene represented in this entry] | Osteogenesis imperfecta 4 (OI4) : An autosomal dominant form of osteogenesis imperfecta, a disorder of bone formation characterized by low bone mass, bone fragility and susceptibility to fractures after minimal trauma. Disease severity ranges from very mild forms without fractures to intrauterine fractures and perinatal lethality. Extraskeletal manifestations, which affect a variable number of patients, are dentinogenesis imperfecta, hearing loss, and blue sclerae. OI4 is characterized by moderately short stature, mild to moderate scoliosis, grayish or white sclera and dentinogenesis imperfecta. [The disease is caused by variants affecting the gene represented in this entry] | Combined osteogenesis imperfecta and Ehlers-Danlos syndrome 1 (OIEDS1) : An autosomal dominant connective tissue disorder characterized by osteopenia, bone fragility, long bone fractures, blue sclerae, joint hyperextensibility, soft and hyperextensible skin, abnormal wound healing, easy bruising, and vascular fragility. [The disease is caused by variants affecting the gene represented in this entry] | Osteoporosis (OSTEOP) : A systemic skeletal disorder characterized by decreased bone mass and deterioration of bone microarchitecture without alteration in the composition of bone. The result is fragile bones and an increased risk of fractures, even after minimal trauma. Osteoporosis is a chronic condition of multifactorial etiology and is usually clinically silent until a fracture occurs. [Disease susceptibility is associated with variants affecting the gene represented in this entry] | [A chromosomal aberration involving COL1A1 is found in dermatofibrosarcoma protuberans. Translocation t(17;22)(q22;q13) with PDGF]

Product Specifications

Recombinant Human Collagen alpha-1 (I) chain (COL1A1), partial is a recombinant protein from Homo sapiens (Human), expressed in Yeast, covering amino acids Confidential, with Confidential tag, molecular weight 47.0kDa, purity Greater than 90% as determined by SDS-PAGE.. Suitable for ELISA and Western Blot applications.
Recommended applications depend on the target, product format, and product-specific validation data. Refer to the product datasheet or Certificate of Analysis (CoA), or contact BioCrest Sci for application guidance.

Biological Activity: Please contact us to obtain bioactivity data.

Storage: Store at -20°C/-80°C upon receipt, aliquoting is necessary for mutiple use. Avoid repeated freeze-thaw cycles.

Shipping & Lead Time: Availability, shipping conditions, and estimated lead time are confirmed in the formal quotation or order confirmation based on the product format, destination, and order requirements.

Protein Biology

Function

Type I collagen is a member of group I collagen (fibrillar forming collagen)

Subcellular Location

Secreted, extracellular space, extracellular matrix

Disease Association

Caffey disease (CAFYD) : An autosomal dominant disorder characterized by an infantile episode of massive subperiosteal new bone formation that typically involves the diaphyses of the long bones, mandible, and clavicles. The involved bones may also appear inflamed, with painful swelling and systemic fever often accompanying the illness. The bone changes usually begin before 5 months of age and resolve before 2 years of age. [The disease is caused by variants affecting the gene represented in this entry] | Ehlers-Danlos syndrome, classic type, 1 (EDSCL1) : A form of Ehlers-Danlos syndrome, a group of connective tissue disorders characterized by skin hyperextensibility, articular hypermobility, and tissue fragility. The main features of classic Ehlers-Danlos syndrome are joint hypermobility and dislocation, and fragile, bruisable skin. EDSCL1 inheritance is autosomal dominant. [The disease may be caused by variants affecting the gene represented in this entry] | Ehlers-Danlos syndrome, arthrochalasia type, 1 (EDSARTH1) : A form of Ehlers-Danlos syndrome, a connective tissue disorder characterized by hyperextensible skin, atrophic cutaneous scars due to tissue fragility and joint hyperlaxity. EDSARTH1 is an autosomal dominant form characterized by frequent congenital hip dislocation and extreme joint laxity with recurrent joint subluxations and minimal skin involvement. [The disease is caused by variants affecting the gene represented in this entry] | Osteogenesis imperfecta 1 (OI1) : An autosomal dominant form of osteogenesis imperfecta, a disorder of bone formation characterized by bone low bone mass, bone fragility and susceptibility to fractures after minimal trauma. Disease severity ranges from very mild forms without fractures to intrauterine fractures and perinatal lethality. Extraskeletal manifestations, which affect a variable number of patients, are dentinogenesis imperfecta, hearing loss, and blue sclerae. OI1 is a non-deforming form with normal height or mild short stature, and no dentinogenesis imperfecta. [The disease is caused by variants affecting the gene represented in this entry] | Osteogenesis imperfecta 2 (OI2) : An autosomal dominant form of osteogenesis imperfecta, a disorder of bone formation characterized by low bone mass, bone fragility and susceptibility to fractures after minimal trauma. Disease severity ranges from very mild forms without fractures to intrauterine fractures and perinatal lethality. Extraskeletal manifestations, which affect a variable number of patients, are dentinogenesis imperfecta, hearing loss, and blue sclerae. OI2 is characterized by bone fragility, with many perinatal fractures, severe bowing of long bones, undermineralization, and death in the perinatal period due to respiratory insufficiency. [The disease is caused by variants affecting the gene represented in this entry] | Osteogenesis imperfecta 3 (OI3) : An autosomal dominant form of osteogenesis imperfecta, a disorder of bone formation characterized by low bone mass, bone fragility and susceptibility to fractures after minimal trauma. Disease severity ranges from very mild forms without fractures to intrauterine fractures and perinatal lethality. Extraskeletal manifestations, which affect a variable number of patients, are dentinogenesis imperfecta, hearing loss, and blue sclerae. OI3 is characterized by progressively deforming bones, very short stature, a triangular face, severe scoliosis, grayish sclera and dentinogenesis imperfecta. [The disease is caused by variants affecting the gene represented in this entry] | Osteogenesis imperfecta 4 (OI4) : An autosomal dominant form of osteogenesis imperfecta, a disorder of bone formation characterized by low bone mass, bone fragility and susceptibility to fractures after minimal trauma. Disease severity ranges from very mild forms without fractures to intrauterine fractures and perinatal lethality. Extraskeletal manifestations, which affect a variable number of patients, are dentinogenesis imperfecta, hearing loss, and blue sclerae. OI4 is characterized by moderately short stature, mild to moderate scoliosis, grayish or white sclera and dentinogenesis imperfecta. [The disease is caused by variants affecting the gene represented in this entry] | Combined osteogenesis imperfecta and Ehlers-Danlos syndrome 1 (OIEDS1) : An autosomal dominant connective tissue disorder characterized by osteopenia, bone fragility, long bone fractures, blue sclerae, joint hyperextensibility, soft and hyperextensible skin, abnormal wound healing, easy bruising, and vascular fragility. [The disease is caused by variants affecting the gene represented in this entry] | Osteoporosis (OSTEOP) : A systemic skeletal disorder characterized by decreased bone mass and deterioration of bone microarchitecture without alteration in the composition of bone. The result is fragile bones and an increased risk of fractures, even after minimal trauma. Osteoporosis is a chronic condition of multifactorial etiology and is usually clinically silent until a fracture occurs. [Disease susceptibility is associated with variants affecting the gene represented in this entry] | [A chromosomal aberration involving COL1A1 is found in dermatofibrosarcoma protuberans. Translocation t(17;22)(q22;q13) with PDGF]

Tissue Specificity

Forms the fibrils of tendon, ligaments and bones. In bones the fibrils are mineralized with calcium hydroxyapatite

Subunit

Trimers of one alpha 2(I) and two alpha 1(I) chains. Interacts with MRC2 (By similarity). Interacts with TRAM2 (PubMed:14749390). Interacts with MFAP4 in a Ca (2+)-dependent manner (By similarity)

Gene: COL1A1  |  Organism: Homo sapiens  |  Synonyms: Alpha-1 type I collagen
Key Publications

Frequently Asked Questions

How do I order or inquire about this product?

Submit the Online Inquiry form with your required quantity and specifications, or email sales@biocrestsci.com. Available sizes, pricing, stock status, documentation, and estimated lead time are confirmed in the formal quotation.

What is the shipping and delivery time?

Availability, dispatch timing, shipping conditions, and delivery estimates depend on the product format, destination, and order requirements. These details are confirmed in the formal quotation or order confirmation.

How should this product be stored?

Follow the storage and handling instructions in the product-specific datasheet and Certificate of Analysis (CoA). Requirements may vary by product form and formulation; contact BioCrest Sci if product-specific guidance is needed.

What quality controls are performed on your products?

Quality-control data such as identity, purity, endotoxin level, or biological activity are provided only when specified on this product page, the datasheet, or Certificate of Analysis (CoA). Biological activity is not implied unless explicitly stated.

Shopping Cart
Scroll to Top