Recombinant Human KeRatin, type I cytoskeletal 14 (KRT14)

Recombinant Human KeRatin, type I cytoskeletal 14 (KRT14) — Structural component of intermediate filaments in basal keratinocytes of stratified epithelia. Purity >90%.

SKU: BCRECP-001026 Category:

Product Specifications

Product SkuBCRECP-001026
Product DescriptionRecombinant Human KeRatin, type I cytoskeletal 14 (KRT14) Protein is expressed from Yeast with N-terminal 6xHis-tagged. It contains 1-472aa. [Accession | P02533].
Uniprot No.P02533
Gene NamesKRT14
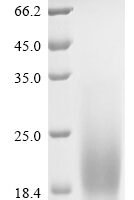
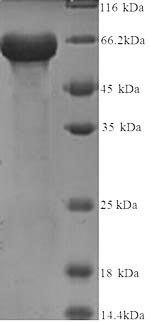
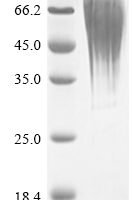
PurityGreater than 90% as determined by SDS-PAGE.
Expression SystemYeast
Expression Region1-472aa
SpeciesHomo sapiens (Human)
Tag InfoN-terminal 6xHis-tagged
Molecular weight53.6kDa
ActivityPlease contact us to obtain bioactivity data.
BufferIf the delivery form is liquid, the default storage buffer is Tris/PBS-based buffer, 5%-50% glycerol. If the delivery form is lyophilized powder, the buffer before lyophilization is Tris/PBS-based buffer, 6% Trehalose.
StorageStore at -20°C/-80°C upon receipt, aliquoting is necessary for mutiple use. Avoid repeated freeze-thaw cycles.
Research AreasSignal Transduction

Function

Structural component of intermediate filaments in basal keratinocytes of stratified epithelia. Forms obligate heteropolymers with the type II keratin KRT5, contributing to the keratin intermediate filament network that confers mechanical strength and structural integrity to the basal layer of the epidermis and other stratified epithelial. The nonhelical tail domain is involved in promoting KRT5-KRT14 filaments to self-organize into large bundles and enhances the mechanical properties involved in resilience of keratin intermediate filaments in vitro.

Biological Context

Subcellular Location: Cytoplasm; Nucleus
Tissue Specificity: Expressed in the corneal epithelium (at protein level). Detected in the basal layer, lowered within the more apically located layers specifically in the stratum spinosum, stratum granulosum but is not detected in stratum corneum. Strongly expressed in the outer root sheath of anagen follicles but not in the germinative matrix, inner root sheath or hair. Found in keratinocytes surrounding the club hair during telogen
Disease Association: Epidermolysis bullosa simplex 1A, generalized severe (EBS1A) : A form of epidermolysis bullosa simplex, a group of skin fragility disorders characterized by skin blistering due to cleavage within the basal layer of keratinocytes, and erosions caused by minor mechanical trauma. There is a broad spectrum of clinical severity ranging from minor blistering on the feet, to subtypes with extracutaneous involvement and a lethal outcome. EBS1A is an autosomal dominant form characterized by generalized intraepidermal skin blistering that begins and is very prominent at birth. EBS1A may be life-threatening in the first year of life. Tendency to blistering diminishes in adolescence. [The disease is caused by variants affecting the gene represented in this entry] | Epidermolysis bullosa simplex 1C, localized (EBS1C) : A form of epidermolysis bullosa simplex, a group of skin fragility disorders characterized by skin blistering due to cleavage within the basal layer of keratinocytes, and erosions caused by minor mechanical trauma. There is a broad spectrum of clinical severity ranging from minor blistering on the feet, to subtypes with extracutaneous involvement and a lethal outcome. EBS1C is an autosomal dominant form with intraepidermal blistering mainly restricted to hands and feet beginning in infancy. Nails may be thick and dystrophic. [The disease is caused by variants affecting the gene represented in this entry] | Epidermolysis bullosa simplex 1B, generalized intermediate (EBS1B) : A form of epidermolysis bullosa simplex, a group of skin fragility disorders characterized by skin blistering due to cleavage within the basal layer of keratinocytes, and erosions caused by minor mechanical trauma. There is a broad spectrum of clinical severity ranging from minor blistering on the feet, to subtypes with extracutaneous involvement and a lethal outcome. EBS1B is an autosomal dominant form characterized by generalized intraepidermal blistering beginning at birth. The tendency to blistering diminishes in adolescence, when it may become localized to hands and feet. [The disease is caused by variants affecting the gene represented in this entry] | Epidermolysis bullosa simplex 1D, generalized, intermediate or severe, autosomal recessive (EBS1D) : A form of epidermolysis bullosa simplex, a group of skin fragility disorders characterized by skin blistering due to cleavage within the basal layer of keratinocytes, and erosions caused by minor mechanical trauma. There is a broad spectrum of clinical severity ranging from minor blistering on the feet, to subtypes with extracutaneous involvement and a lethal outcome. EBS1D is an autosomal recessive form characterized by blistering beginning at birth or early childhood. In some patients hands and feet are primarily affected, and in others blistering anywhere on the body may occur. In some patients the condition improves with age. [The disease is caused by variants affecting the gene represented in this entry] | Naegeli-Franceschetti-Jadassohn syndrome (NFJS) : A rare autosomal dominant form of ectodermal dysplasia. The cardinal features are absence of dermatoglyphics (fingerprints), reticular cutaneous hyperpigmentation (starting at about the age of 2 years without a preceding inflammatory stage), palmoplantar keratoderma, hypohidrosis with diminished sweat gland function and discomfort provoked by heat, nail dystrophy, and tooth enamel defects. [The disease is caused by variants affecting the gene represented in this entry] | Dermatopathia pigmentosa reticularis (DPR) : A rare ectodermal dysplasia characterized by lifelong persistent reticulate hyperpigmentation, non-cicatricial alopecia, and nail dystrophy. Variable features include adermatoglyphia, hypohidrosis or hyperhidrosis, and palmoplantar hyperkeratosis. [The disease is caused by variants affecting the gene represented in this entry]

Product Specifications

Recombinant Human KeRatin, type I cytoskeletal 14 (KRT14) is a recombinant protein from Homo sapiens (Human), expressed in Yeast, covering amino acids 1-472aa, with N-terminal 6xHis-tagged tag, molecular weight 53.6kDa, purity Greater than 90% as determined by SDS-PAGE.. Suitable for ELISA and Western Blot applications.

Recommended applications depend on the target, product format, and product-specific validation data. Refer to the product datasheet or Certificate of Analysis (CoA), or contact BioCrest Sci for application guidance.

Biological Activity: Please contact us to obtain bioactivity data.

Storage: Store at -20°C/-80°C upon receipt, aliquoting is necessary for mutiple use. Avoid repeated freeze-thaw cycles.

Shipping & Lead Time: Availability, shipping conditions, and estimated lead time are confirmed in the formal quotation or order confirmation based on the product format, destination, and order requirements.

Protein Biology

Function

Structural component of intermediate filaments in basal keratinocytes of stratified epithelia. Forms obligate heteropolymers with the type II keratin KRT5, contributing to the keratin intermediate filament network that confers mechanical strength and structural integrity to the basal layer of the epidermis and other stratified epithelial (PubMed:11724817, PubMed:1694855, PubMed:1720261). The nonhelical tail domain is involved in promoting KRT5-KRT14 filaments to self-organize into large bundles and enhances the mechanical properties involved in resilience of keratin intermediate filaments in vitro (PubMed:11724817)

Subcellular Location

Cytoplasm; Nucleus

Disease Association

Epidermolysis bullosa simplex 1A, generalized severe (EBS1A) : A form of epidermolysis bullosa simplex, a group of skin fragility disorders characterized by skin blistering due to cleavage within the basal layer of keratinocytes, and erosions caused by minor mechanical trauma. There is a broad spectrum of clinical severity ranging from minor blistering on the feet, to subtypes with extracutaneous involvement and a lethal outcome. EBS1A is an autosomal dominant form characterized by generalized intraepidermal skin blistering that begins and is very prominent at birth. EBS1A may be life-threatening in the first year of life. Tendency to blistering diminishes in adolescence. [The disease is caused by variants affecting the gene represented in this entry] | Epidermolysis bullosa simplex 1C, localized (EBS1C) : A form of epidermolysis bullosa simplex, a group of skin fragility disorders characterized by skin blistering due to cleavage within the basal layer of keratinocytes, and erosions caused by minor mechanical trauma. There is a broad spectrum of clinical severity ranging from minor blistering on the feet, to subtypes with extracutaneous involvement and a lethal outcome. EBS1C is an autosomal dominant form with intraepidermal blistering mainly restricted to hands and feet beginning in infancy. Nails may be thick and dystrophic. [The disease is caused by variants affecting the gene represented in this entry] | Epidermolysis bullosa simplex 1B, generalized intermediate (EBS1B) : A form of epidermolysis bullosa simplex, a group of skin fragility disorders characterized by skin blistering due to cleavage within the basal layer of keratinocytes, and erosions caused by minor mechanical trauma. There is a broad spectrum of clinical severity ranging from minor blistering on the feet, to subtypes with extracutaneous involvement and a lethal outcome. EBS1B is an autosomal dominant form characterized by generalized intraepidermal blistering beginning at birth. The tendency to blistering diminishes in adolescence, when it may become localized to hands and feet. [The disease is caused by variants affecting the gene represented in this entry] | Epidermolysis bullosa simplex 1D, generalized, intermediate or severe, autosomal recessive (EBS1D) : A form of epidermolysis bullosa simplex, a group of skin fragility disorders characterized by skin blistering due to cleavage within the basal layer of keratinocytes, and erosions caused by minor mechanical trauma. There is a broad spectrum of clinical severity ranging from minor blistering on the feet, to subtypes with extracutaneous involvement and a lethal outcome. EBS1D is an autosomal recessive form characterized by blistering beginning at birth or early childhood. In some patients hands and feet are primarily affected, and in others blistering anywhere on the body may occur. In some patients the condition improves with age. [The disease is caused by variants affecting the gene represented in this entry] | Naegeli-Franceschetti-Jadassohn syndrome (NFJS) : A rare autosomal dominant form of ectodermal dysplasia. The cardinal features are absence of dermatoglyphics (fingerprints), reticular cutaneous hyperpigmentation (starting at about the age of 2 years without a preceding inflammatory stage), palmoplantar keratoderma, hypohidrosis with diminished sweat gland function and discomfort provoked by heat, nail dystrophy, and tooth enamel defects. [The disease is caused by variants affecting the gene represented in this entry] | Dermatopathia pigmentosa reticularis (DPR) : A rare ectodermal dysplasia characterized by lifelong persistent reticulate hyperpigmentation, non-cicatricial alopecia, and nail dystrophy. Variable features include adermatoglyphia, hypohidrosis or hyperhidrosis, and palmoplantar hyperkeratosis. [The disease is caused by variants affecting the gene represented in this entry]

Tissue Specificity

Expressed in the corneal epithelium (at protein level) (PubMed:26758872). Detected in the basal layer, lowered within the more apically located layers specifically in the stratum spinosum, stratum granulosum but is not detected in stratum corneum. Strongly expressed in the outer root sheath of anagen follicles but not in the germinative matrix, inner root sheath or hair (PubMed:9457912). Found in keratinocytes surrounding the club hair during telogen (PubMed:9457912)

Subunit

Heterodimers composed of type I and type II keratin; forms parallel coiled-coil heterodimers (PubMed:11724817, PubMed:1694855, PubMed:22705788, PubMed:24940650). Two heterodimers associate in an antiparallel manner to form heterotetramers, which further assemble into higher-order intermediate filaments (PubMed:1694855). Forms a disulfide-linked heterodimer (via 2B domains) with KRT5 (via 2B domains) (PubMed:1694855, PubMed:22705788, PubMed:24940650). Forms a heterodimer with KRT1; the interaction is more abundant in the absence of KRT5 (By similarity). Interacts with PLEC isoform 1C, when in a heterodimer with KRT5 (PubMed:24940650). Interacts with TRADD and with keratin filaments (PubMed:11684708). Interacts with EPPK1 (By similarity). Interacts with KLHL24 (PubMed:27798626). Interacts with PKP1 (via N-terminus) and PKP2 (PubMed:10852826)

Gene: KRT14  |  Organism: Homo sapiens  |  Synonyms: Cytokeratin-14; Keratin-14
Key Publications

Frequently Asked Questions

How do I order or inquire about this product?

Submit the Online Inquiry form with your required quantity and specifications, or email sales@biocrestsci.com. Available sizes, pricing, stock status, documentation, and estimated lead time are confirmed in the formal quotation.

What is the shipping and delivery time?

Availability, dispatch timing, shipping conditions, and delivery estimates depend on the product format, destination, and order requirements. These details are confirmed in the formal quotation or order confirmation.

How should this product be stored?

Follow the storage and handling instructions in the product-specific datasheet and Certificate of Analysis (CoA). Requirements may vary by product form and formulation; contact BioCrest Sci if product-specific guidance is needed.

What quality controls are performed on your products?

Quality-control data such as identity, purity, endotoxin level, or biological activity are provided only when specified on this product page, the datasheet, or Certificate of Analysis (CoA). Biological activity is not implied unless explicitly stated.

Shopping Cart
Scroll to Top