,

Recombinant Human Succinate dehydrogenase [ubiquinone] flavoprotein subunit, mitochondrial (SDHA), partial

Recombinant Human Succinate dehydrogenase [ubiquinone] flavoprotein subunit, mitochondrial (SDHA), partial — Flavoprotein (FP) subunit of succinate dehydrogenase (SDH) that is involved in complex II of the mitochondrial electron transport chain and is responsible for transferring electrons from s…

SKU: BCRECP-001595 Categories: ,

Product Specifications

Product SkuBCRECP-001595
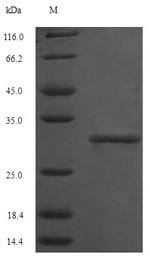
Product DescriptionRecombinant Human Succinate dehydrogenase [ubiquinone] flavoprotein subunit, mitochondrial (SDHA) Protein is expressed from E.coli with N-terminal 6xHis-tagged. It contains 44-293aa. [Accession | P31040].
Uniprot No.P31040
Gene NamesSDHA
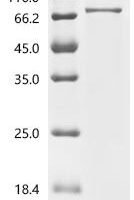
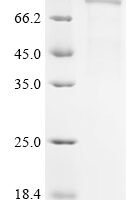
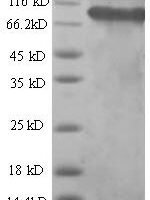
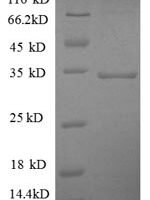
PurityGreater than 90% as determined by SDS-PAGE.
Expression SystemE.coli
Expression Region44-293aa
SpeciesHomo sapiens (Human)
Tag InfoN-terminal 6xHis-tagged
Molecular weight31.1kDa
ActivityPlease contact us to obtain bioactivity data.
BufferIf the delivery form is liquid, the default storage buffer is Tris/PBS-based buffer, 5%-50% glycerol. If the delivery form is lyophilized powder, the buffer before lyophilization is Tris/PBS-based buffer, 6% Trehalose.
StorageStore at -20°C/-80°C upon receipt, aliquoting is necessary for mutiple use. Avoid repeated freeze-thaw cycles.
Research AreasTransport

Function

Flavoprotein (FP) subunit of succinate dehydrogenase (SDH) that is involved in complex II of the mitochondrial electron transport chain and is responsible for transferring electrons from succinate to ubiquinone (coenzyme Q). SDH also oxidizes malate to the non-canonical enol form of oxaloacetate, enol-oxaloacetate. Enol-oxaloacetate, which is a potent inhibitor of the succinate dehydrogenase activity, is further isomerized into keto-oxaloacetate.

Biological Context

Subcellular Location: Mitochondrion inner membrane (Peripheral membrane protein)
Disease Association: Mitochondrial complex II deficiency, nuclear type 1 (MC2DN1) : A disorder of the mitochondrial respiratory chain with heterogeneous clinical manifestations. Clinical features include psychomotor regression in infants, poor growth with lack of speech development, severe spastic quadriplegia, dystonia, progressive leukoencephalopathy, muscle weakness, exercise intolerance, cardiomyopathy. Some patients manifest Leigh syndrome or Kearns-Sayre syndrome. MC2DN1 inheritance is autosomal recessive. [The disease is caused by variants affecting the gene represented in this entry] | Leigh syndrome (LS) : An early-onset progressive neurodegenerative disorder characterized by the presence of focal, bilateral lesions in one or more areas of the central nervous system including the brainstem, thalamus, basal ganglia, cerebellum and spinal cord. Clinical features depend on which areas of the central nervous system are involved and include subacute onset of psychomotor retardation, hypotonia, ataxia, weakness, vision loss, eye movement abnormalities, seizures, and dysphagia. [The disease is caused by variants affecting the gene represented in this entry] | Cardiomyopathy, dilated, 1GG (CMD1GG) : A disorder characterized by ventricular dilation and impaired systolic function, resulting in congestive heart failure and arrhythmia. Patients are at risk of premature death. [The disease is caused by variants affecting the gene represented in this entry] | Pheochromocytoma/paraganglioma syndrome 5 (PPGL5) : A form of pheochromocytoma/paraganglioma syndrome, a tumor predisposition syndrome characterized by the development of neuroendocrine tumors, usually in adulthood. Pheochromocytomas are catecholamine-producing tumors that arise from chromaffin cells in the adrenal medulla. Paragangliomas develop from sympathetic paraganglia in the thorax, abdomen, and pelvis, as well as from parasympathetic paraganglia in the head and neck. PPGL5 inheritance is autosomal dominant. [The disease is caused by variants affecting the gene represented in this entry] | Neurodegeneration with ataxia and late-onset optic atrophy (NDAXOA) : An autosomal dominant disorder characterized by slowly progressive cerebellar and gait ataxia, optic atrophy, and myopathy or myalgia. Additional features can include cardiomyopathy, psychiatric disturbances, and peripheral sensory impairment. Disease onset is usually in mid-adulthood. [The disease is caused by variants affecting the gene represented in this entry]
Pathway: Carbohydrate metabolism; tricarboxylic acid cycle; fumarate from succinate (eukaryal route): step 1/1

Product Specifications

Recombinant Human Succinate dehydrogenase [ubiquinone] flavoprotein subunit, mitochondrial (SDHA), partial is a recombinant protein from Homo sapiens (Human), expressed in E.coli, covering amino acids 44-293aa, with N-terminal 6xHis-tagged tag, molecular weight 31.1kDa, purity Greater than 90% as determined by SDS-PAGE.. Suitable for ELISA and Western Blot applications.

Recommended applications depend on the target, product format, and product-specific validation data. Refer to the product datasheet or Certificate of Analysis (CoA), or contact BioCrest Sci for application guidance.

Biological Activity: Please contact us to obtain bioactivity data.

Storage: Store at -20°C/-80°C upon receipt, aliquoting is necessary for mutiple use. Avoid repeated freeze-thaw cycles.

Shipping & Lead Time: Availability, shipping conditions, and estimated lead time are confirmed in the formal quotation or order confirmation based on the product format, destination, and order requirements.

Protein Biology

Function

Flavoprotein (FP) subunit of succinate dehydrogenase (SDH) that is involved in complex II of the mitochondrial electron transport chain and is responsible for transferring electrons from succinate to ubiquinone (coenzyme Q) (PubMed:10746566, PubMed:24781757). SDH also oxidizes malate to the non-canonical enol form of oxaloacetate, enol-oxaloacetate (By similarity). Enol-oxaloacetate, which is a potent inhibitor of the succinate dehydrogenase activity, is further isomerized into keto-oxaloacetate (By similarity). Can act as a tumor suppressor (PubMed:20484225)

Subcellular Location

Mitochondrion inner membrane (Peripheral membrane protein)

Disease Association

Mitochondrial complex II deficiency, nuclear type 1 (MC2DN1) : A disorder of the mitochondrial respiratory chain with heterogeneous clinical manifestations. Clinical features include psychomotor regression in infants, poor growth with lack of speech development, severe spastic quadriplegia, dystonia, progressive leukoencephalopathy, muscle weakness, exercise intolerance, cardiomyopathy. Some patients manifest Leigh syndrome or Kearns-Sayre syndrome. MC2DN1 inheritance is autosomal recessive. [The disease is caused by variants affecting the gene represented in this entry] | Leigh syndrome (LS) : An early-onset progressive neurodegenerative disorder characterized by the presence of focal, bilateral lesions in one or more areas of the central nervous system including the brainstem, thalamus, basal ganglia, cerebellum and spinal cord. Clinical features depend on which areas of the central nervous system are involved and include subacute onset of psychomotor retardation, hypotonia, ataxia, weakness, vision loss, eye movement abnormalities, seizures, and dysphagia. [The disease is caused by variants affecting the gene represented in this entry] | Cardiomyopathy, dilated, 1GG (CMD1GG) : A disorder characterized by ventricular dilation and impaired systolic function, resulting in congestive heart failure and arrhythmia. Patients are at risk of premature death. [The disease is caused by variants affecting the gene represented in this entry] | Pheochromocytoma/paraganglioma syndrome 5 (PPGL5) : A form of pheochromocytoma/paraganglioma syndrome, a tumor predisposition syndrome characterized by the development of neuroendocrine tumors, usually in adulthood. Pheochromocytomas are catecholamine-producing tumors that arise from chromaffin cells in the adrenal medulla. Paragangliomas develop from sympathetic paraganglia in the thorax, abdomen, and pelvis, as well as from parasympathetic paraganglia in the head and neck. PPGL5 inheritance is autosomal dominant. [The disease is caused by variants affecting the gene represented in this entry] | Neurodegeneration with ataxia and late-onset optic atrophy (NDAXOA) : An autosomal dominant disorder characterized by slowly progressive cerebellar and gait ataxia, optic atrophy, and myopathy or myalgia. Additional features can include cardiomyopathy, psychiatric disturbances, and peripheral sensory impairment. Disease onset is usually in mid-adulthood. [The disease is caused by variants affecting the gene represented in this entry]

Subunit

Component of complex II composed of four subunits: the flavoprotein (FP) SDHA, iron-sulfur protein (IP) SDHB, and a cytochrome b560 composed of SDHC and SDHD (PubMed:37098072). Interacts with SDHAF2/SDH5; interaction is required for FAD attachment (PubMed:19628817, PubMed:32887801). Interacts with TRAP1 (PubMed:23747254). Interacts with LACC1 (PubMed:28593945)

Pathway

Carbohydrate metabolism; tricarboxylic acid cycle; fumarate from succinate (eukaryal route): step 1/1

Gene: SDHA  |  Organism: Homo sapiens  |  Synonyms: Flavoprotein subunit of complex II; Malate dehydrogenase [quinone] flavoprotein subunit
Key Publications

Frequently Asked Questions

How do I order or inquire about this product?

Submit the Online Inquiry form with your required quantity and specifications, or email sales@biocrestsci.com. Available sizes, pricing, stock status, documentation, and estimated lead time are confirmed in the formal quotation.

What is the shipping and delivery time?

Availability, dispatch timing, shipping conditions, and delivery estimates depend on the product format, destination, and order requirements. These details are confirmed in the formal quotation or order confirmation.

How should this product be stored?

Follow the storage and handling instructions in the product-specific datasheet and Certificate of Analysis (CoA). Requirements may vary by product form and formulation; contact BioCrest Sci if product-specific guidance is needed.

What quality controls are performed on your products?

Quality-control data such as identity, purity, endotoxin level, or biological activity are provided only when specified on this product page, the datasheet, or Certificate of Analysis (CoA). Biological activity is not implied unless explicitly stated.

Shopping Cart
Scroll to Top