,

Recombinant Human Carnitine O-palmitoyltransferase 2, mitochondrial (CPT2), partial

Recombinant Human Carnitine O-palmitoyltransferase 2, mitochondrial (CPT2), partial — Involved in the intramitochondrial synthesis of acylcarnitines from accumulated acyl-CoA metabolites. Purity >90%.

SKU: BCRECP-000423 Categories: ,

Product Specifications

Product SkuBCRECP-000423
Product DescriptionRecombinant Human Carnitine O-palmitoyltransferase 2, mitochondrial (CPT2) Protein is expressed from E.coli with N-terminal 10xHis-tagged and C-terminal Myc-tagged. It contains 209-658aa. [Accession | P23786].
Uniprot No.P23786
Gene NamesCPT2
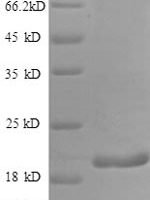
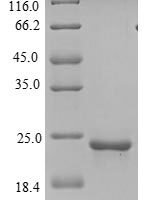
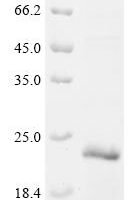
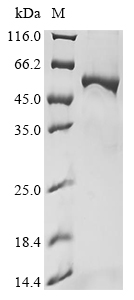
PurityGreater than 90% as determined by SDS-PAGE.
Expression SystemE.coli
Expression Region209-658aa
SpeciesHomo sapiens (Human)
Tag InfoN-terminal 10xHis-tagged and C-terminal Myc-tagged
Molecular weight57.6kDa
ActivityPlease contact us to obtain bioactivity data.
BufferIf the delivery form is liquid, the default storage buffer is Tris/PBS-based buffer, 5%-50% glycerol. If the delivery form is lyophilized powder, the buffer before lyophilization is Tris/PBS-based buffer, 6% Trehalose.
StorageStore at -20°C/-80°C upon receipt, aliquoting is necessary for mutiple use. Avoid repeated freeze-thaw cycles.
Research AreasCancer

Function

Involved in the intramitochondrial synthesis of acylcarnitines from accumulated acyl-CoA metabolites. Reconverts acylcarnitines back into the respective acyl-CoA esters that can then undergo beta-oxidation, an essential step for the mitochondrial uptake of long-chain fatty acids and their subsequent beta-oxidation in the mitochondrion. Active with medium (C8-C12) and long-chain (C14-C18) acyl-CoA esters.

Biological Context

Subcellular Location: Mitochondrion inner membrane (Peripheral membrane protein)
Disease Association: Carnitine palmitoyltransferase 2 deficiency, myopathic, stress-induced (CPT2D) : An autosomal recessive disorder of mitochondrial long-chain fatty acid oxidation, characterized by recurrent myoglobinuria, episodes of muscle pain, stiffness, and rhabdomyolysis. These symptoms are exacerbated by prolonged exercise, fasting, cold, or viral infection. CPT2DM affects most frequently children or young adults, and severity of attacks is highly variable. Myoglobinuria can cause kidney failure and death. [The disease is caused by variants affecting the gene represented in this entry] | Carnitine palmitoyltransferase 2 deficiency, infantile (CPT2DI) : An autosomal recessive disorder of mitochondrial long-chain fatty acid oxidation, characterized by hepatic or hepato-cardio-muscular manifestations with onset in infancy. Clinical features include hypoketotic hypoglycemia, lethargy, seizures, hepatomegaly, liver dysfunction, cardiomegaly and dilated cardiomyopathy. [The disease is caused by variants affecting the gene represented in this entry] | Carnitine palmitoyltransferase 2 deficiency, lethal neonatal (CPT2DLN) : An autosomal recessive disorder of mitochondrial long-chain fatty acid oxidation with fatal outcome, presenting shortly after birth. It is characterized by respiratory distress, seizures, altered mental status, hepatomegaly, cardiomegaly, cardiac arrhythmia, and, in many cases, dysmorphic features, renal dysgenesis, and migration defects. recessive. [The disease is caused by variants affecting the gene represented in this entry] | Encephalopathy, acute, infection-induced, 4 (IIAE4) : A severe neurologic complication of an infection. It manifests within days in otherwise healthy children after common viral infections, without evidence of viral infection of the brain or inflammatory cell infiltration. In affected children, high-grade fever is accompanied within 12 to 48 hours by febrile convulsions, often leading to coma, multiple-organ failure, brain edema, and high morbidity and mortality. The infections are usually viral, particularly influenza, although other viruses and even mycoplasma have been found to cause the disorder. [Disease susceptibility is associated with variants affecting the gene represented in this entry. CPT2 polymorphic variants do not cause classical carnitine palmitoyltransferase 2 deficiency, and patients harboring any of them are asymptomatic most of the time. However, they are prone to viral infection (high fever)-related encephalopathy ]
Pathway: Lipid metabolism; fatty acid beta-oxidation

Product Specifications

Recombinant Human Carnitine O-palmitoyltransferase 2, mitochondrial (CPT2), partial is a recombinant protein from Homo sapiens (Human), expressed in E.coli, covering amino acids 209-658aa, with N-terminal 10xHis-tagged and C-terminal Myc-tagged tag, molecular weight 57.6kDa, purity Greater than 90% as determined by SDS-PAGE.. Suitable for ELISA and Western Blot applications.

Recommended applications depend on the target, product format, and product-specific validation data. Refer to the product datasheet or Certificate of Analysis (CoA), or contact BioCrest Sci for application guidance.

Biological Activity: Please contact us to obtain bioactivity data.

Storage: Store at -20°C/-80°C upon receipt, aliquoting is necessary for mutiple use. Avoid repeated freeze-thaw cycles.

Shipping & Lead Time: Availability, shipping conditions, and estimated lead time are confirmed in the formal quotation or order confirmation based on the product format, destination, and order requirements.

Protein Biology

Function

Involved in the intramitochondrial synthesis of acylcarnitines from accumulated acyl-CoA metabolites (PubMed:20538056, PubMed:24780397). Reconverts acylcarnitines back into the respective acyl-CoA esters that can then undergo beta-oxidation, an essential step for the mitochondrial uptake of long-chain fatty acids and their subsequent beta-oxidation in the mitochondrion. Active with medium (C8-C12) and long-chain (C14-C18) acyl-CoA esters (PubMed:20538056)

Subcellular Location

Mitochondrion inner membrane (Peripheral membrane protein)

Disease Association

Carnitine palmitoyltransferase 2 deficiency, myopathic, stress-induced (CPT2D) : An autosomal recessive disorder of mitochondrial long-chain fatty acid oxidation, characterized by recurrent myoglobinuria, episodes of muscle pain, stiffness, and rhabdomyolysis. These symptoms are exacerbated by prolonged exercise, fasting, cold, or viral infection. CPT2DM affects most frequently children or young adults, and severity of attacks is highly variable. Myoglobinuria can cause kidney failure and death. [The disease is caused by variants affecting the gene represented in this entry] | Carnitine palmitoyltransferase 2 deficiency, infantile (CPT2DI) : An autosomal recessive disorder of mitochondrial long-chain fatty acid oxidation, characterized by hepatic or hepato-cardio-muscular manifestations with onset in infancy. Clinical features include hypoketotic hypoglycemia, lethargy, seizures, hepatomegaly, liver dysfunction, cardiomegaly and dilated cardiomyopathy. [The disease is caused by variants affecting the gene represented in this entry] | Carnitine palmitoyltransferase 2 deficiency, lethal neonatal (CPT2DLN) : An autosomal recessive disorder of mitochondrial long-chain fatty acid oxidation with fatal outcome, presenting shortly after birth. It is characterized by respiratory distress, seizures, altered mental status, hepatomegaly, cardiomegaly, cardiac arrhythmia, and, in many cases, dysmorphic features, renal dysgenesis, and migration defects. recessive. [The disease is caused by variants affecting the gene represented in this entry] | Encephalopathy, acute, infection-induced, 4 (IIAE4) : A severe neurologic complication of an infection. It manifests within days in otherwise healthy children after common viral infections, without evidence of viral infection of the brain or inflammatory cell infiltration. In affected children, high-grade fever is accompanied within 12 to 48 hours by febrile convulsions, often leading to coma, multiple-organ failure, brain edema, and high morbidity and mortality. The infections are usually viral, particularly influenza, although other viruses and even mycoplasma have been found to cause the disorder. [Disease susceptibility is associated with variants affecting the gene represented in this entry. CPT2 polymorphic variants do not cause classical carnitine palmitoyltransferase 2 deficiency, and patients harboring any of them are asymptomatic most of the time. However, they are prone to viral infection (high fever)-related encephalopathy (PubMed:21697855)]

Pathway

Lipid metabolism; fatty acid beta-oxidation

Gene: CPT2  |  Organism: Homo sapiens  |  Synonyms: Carnitine palmitoyltransferase II
Key Publications

Frequently Asked Questions

How do I order or inquire about this product?

Submit the Online Inquiry form with your required quantity and specifications, or email sales@biocrestsci.com. Available sizes, pricing, stock status, documentation, and estimated lead time are confirmed in the formal quotation.

What is the shipping and delivery time?

Availability, dispatch timing, shipping conditions, and delivery estimates depend on the product format, destination, and order requirements. These details are confirmed in the formal quotation or order confirmation.

How should this product be stored?

Follow the storage and handling instructions in the product-specific datasheet and Certificate of Analysis (CoA). Requirements may vary by product form and formulation; contact BioCrest Sci if product-specific guidance is needed.

What quality controls are performed on your products?

Quality-control data such as identity, purity, endotoxin level, or biological activity are provided only when specified on this product page, the datasheet, or Certificate of Analysis (CoA). Biological activity is not implied unless explicitly stated.

Shopping Cart
Scroll to Top